Chinese Journal of Catalysis
2026, Vol. 91
Online: 18 December 2026

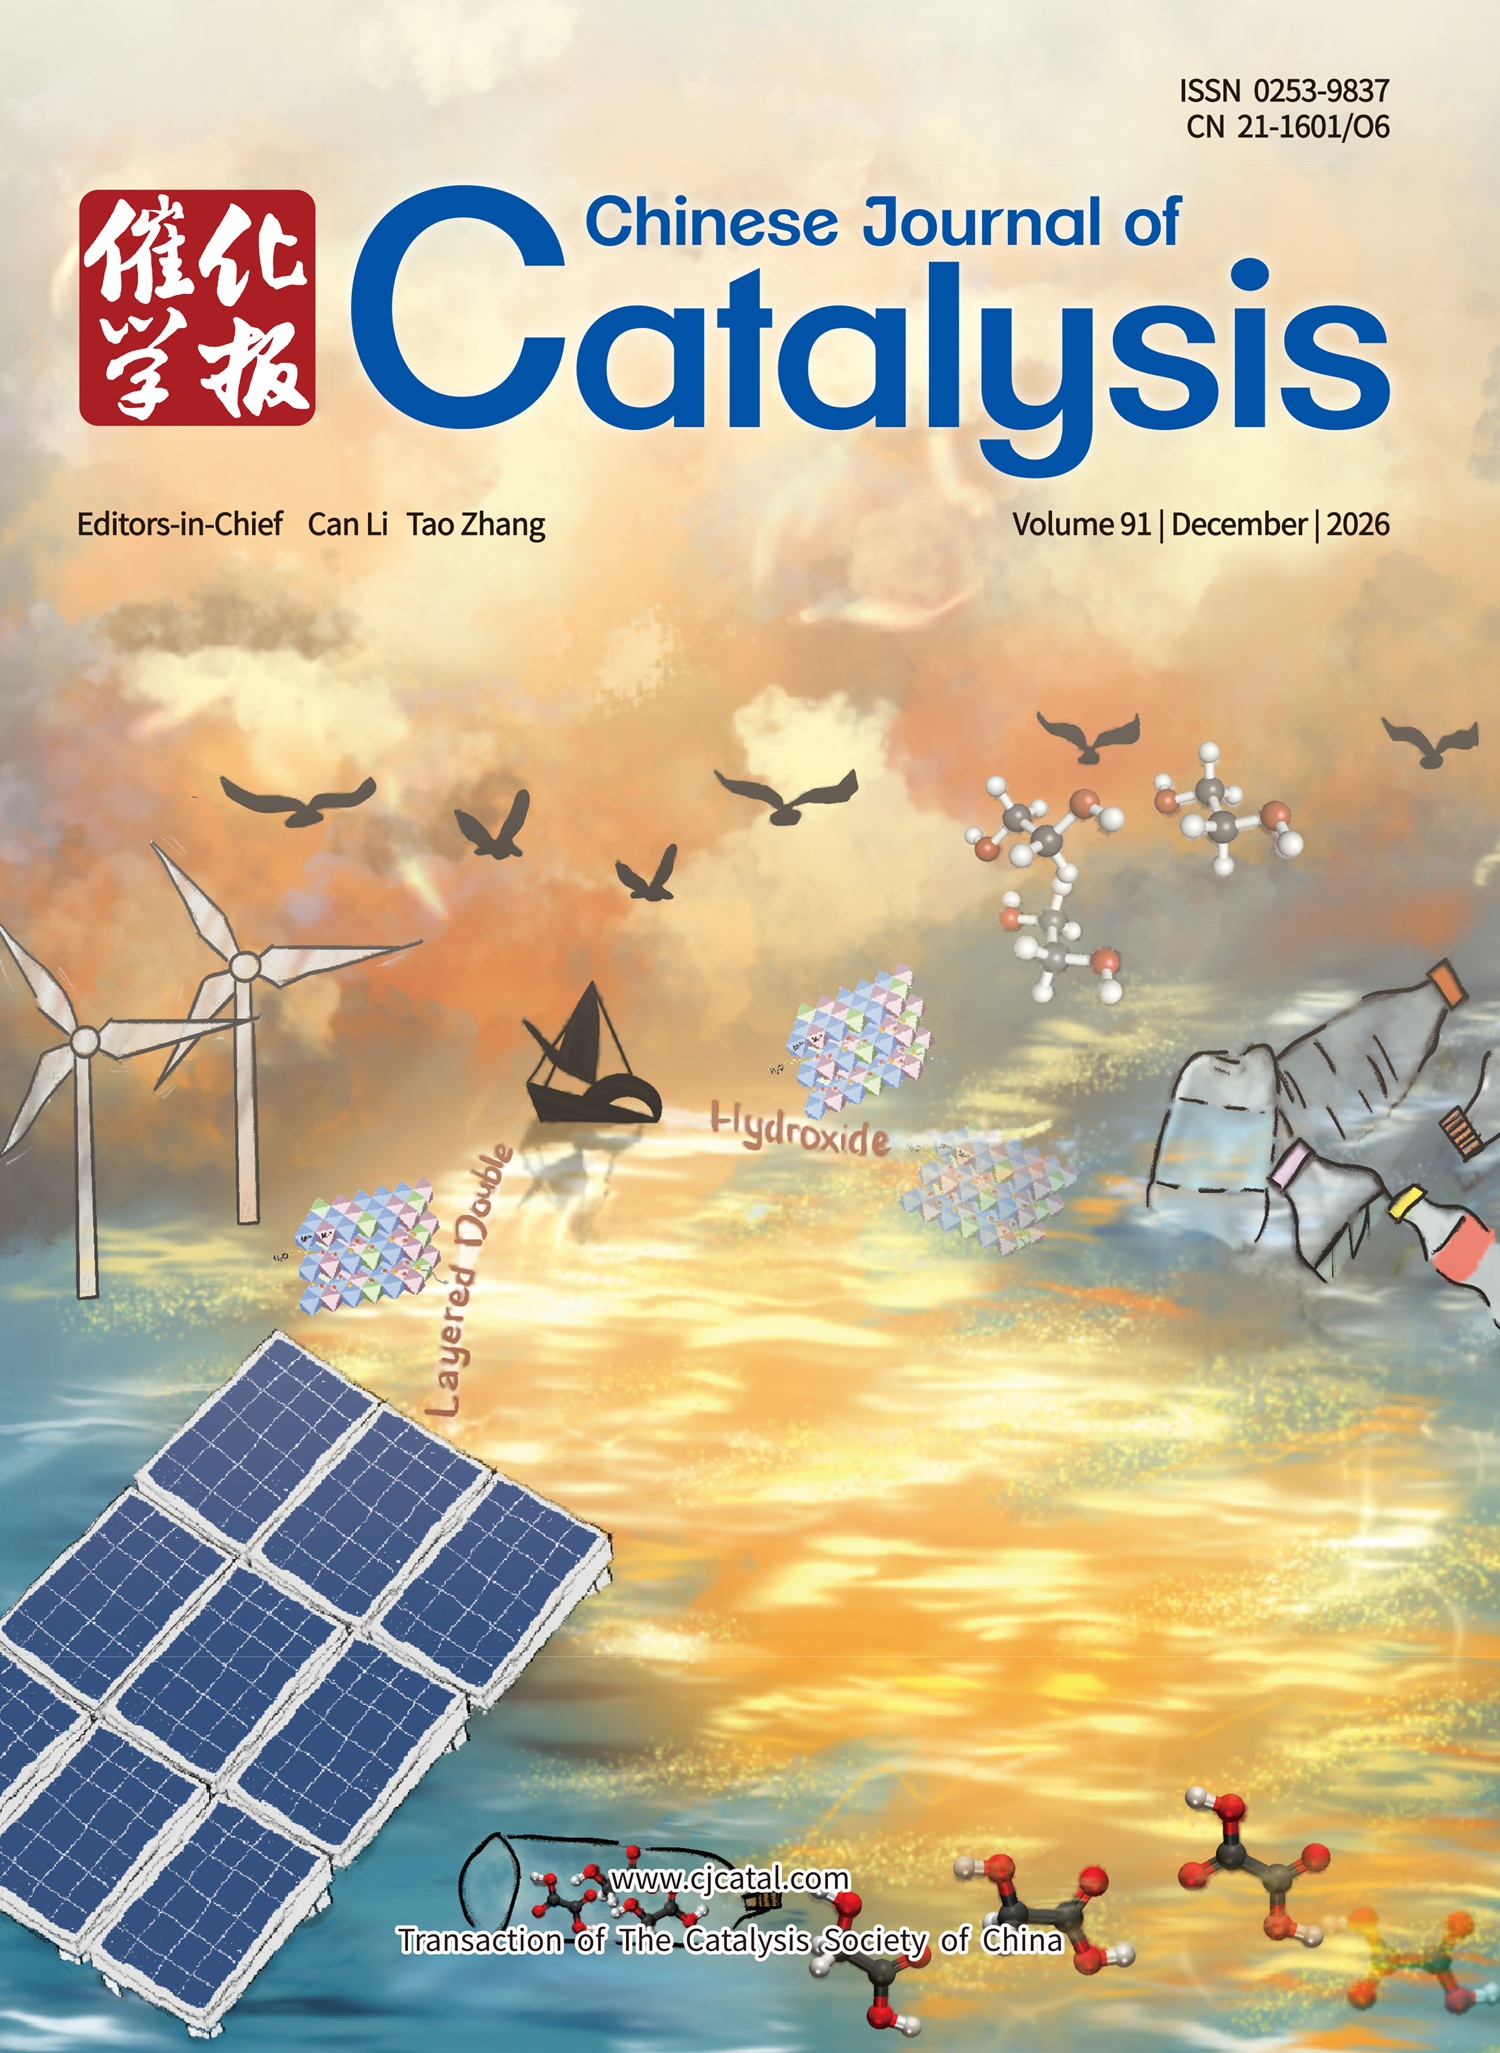
Cover: In their article on pages 103–121, Prof. Wee-Jun Ong, Prof. Xianhai Zeng and coworkers systematically investigated electronic structure modulation in Cu2+-mediated layered double hydroxide catalyst for selective C2 formation via CO2RR and PET upcycling. The optimized electronic structure promotes charge transfer, stabilizes key intermediates, and enhances catalytic selectivity. The cover art depicts a floating solar panel harnessing sunlight to drive the simultaneous electrochemical conversion of CO2 and PET-derived ethylene glycol into value-added C2 products. Floating plastic waste symbolizes circular recycling, while molecular structures represent the products. Renewable energy and natural scenery emphasize a sustainable pathway toward carbon neutrality and a circular chemical economy.
For Selected: Toggle Thumbnails
Reviews
Recent advances in catalytic conversion technologies for plastic waste upgrading
Yang Wan, Bao Zou, Xianghai Song, Yan Liu, Jianmin Luo, Xin Liu, Xin Li, Pengwei Huo
2026, 91:  1-49.  DOI: 10.1016/S1872-2067(26)65230-8
Abstract ( 123 )   HTML ( 4 )   PDF (35000KB) ( 39 )  

The rapid growth in plastic consumption and improper disposal have exacerbated plastic pollution (“white pollution”) and resulted in a significant waste of hydrocarbon resources. Based on the concepts of circular economy and “carbon peaking, carbon neutrality”, the upgrading and transformation of plastic waste is receiving increasing attention. In recent years, catalytic conversion technology has become a key method to solve the contradiction between plastic pollution and resource utilization, including but not limited to photocatalysis, thermocatalysis, electrocatalysis, and biocatalysis. It can not only directly and effectively reduce environmental pollution, but also produce high-value chemicals, fuels from waste resources, thereby alleviating the global energy shortage. Based on this, this review systematically elucidates the direct and indirect pathways for plastic upcycling, and provides an in-depth comparison of the reaction mechanisms, catalyst design strategies, and selective activation rules of chemical bonds across different catalytic approaches. For the first time, it systematically integrates the latest research progress on photocatalytic, thermocatalytic, electrocatalytic, biocatalytic, and synergistic catalytic technologies for the upgrading conversion of typical plastics, clarifies the application scenarios and key bottlenecks of various catalytic methods, reveals the relationship between catalyst microstructure and selective bond cleavage, and proposes important directions for the future design of catalytic systems. This review aims to provide a theoretical framework for the resource-oriented conversion of waste plastics, thereby advancing the development of a green circular economy and supporting sustainable development goals.

CdS-based S-scheme heterojunctions for photocatalytic hydrogen evolution
Haoyu Long, Xinhe Wu, Dmitry Selishchev, Jahan B. Ghasemi, Jiaguo Yu, Huogen Yu
2026, 91:  50-67.  DOI: 10.1016/S1872-2067(26)65210-2
Abstract ( 67 )   HTML ( 5 )   PDF (11253KB) ( 12 )  

Cadmium sulfide (CdS) is a promising visible-light-responsive photocatalyst for solar-driven hydrogen (H2) production owing to its suitable band structure, yet pristine CdS generally exhibits only moderate photocatalytic activity because of rapid charge-carrier recombination and severe photocorrosion. Step-scheme (S-scheme) heterojunctions offer a structural solution to circumvent these bottlenecks. By driving the directional recombination of less reactive charge carriers at the physical interface, the S-scheme mechanism preserves the charge carriers with the strongest redox potentials. This specific pathway maintains robust H2 evolution kinetics while transferring oxidizing holes away from CdS, effectively mitigating self-oxidation. This review systematically summarizes recent advancements in CdS based S-scheme heterojunctions. We elucidate the fundamental thermodynamic principles and evaluate rational interfacial design strategies across diverse structural dimensions. Furthermore, we highlight the rigorous experimental evidence required to verify charge transfer pathways using advanced dynamic characterization techniques. Finally, we outline current challenges and future directions toward practical applications.

Atomically dispersed cocatalysts for photocatalysis: From single and dual atoms to high-entropy alloys
Ting Chen, Siyu Wu, Kangle Lv, Qin Li
2026, 91:  68-92.  DOI: 10.1016/S1872-2067(26)65261-8
Abstract ( 36 )   HTML ( 3 )   PDF (5951KB) ( 10 )  

With increasing demands for energy conversion and environmental remediation, photocatalysis has become an important route for efficient solar-energy utilization and small-molecule conversion owing to its green and renewable characteristics. Atomically dispersed photocatalytic cocatalysts, featuring unique structures and maximized atom utilization, bridge heterogeneous and homogeneous catalysis. In recent years, this field has developed rapidly from single-atom catalysts to synergistic dual-atom sites and further to structurally complex high-entropy alloys. This review systematically summarized recent progress in atomically dispersed cocatalysts from single atoms and dual atoms to high-entropy alloys. Emphasis was placed on their synthetic methodologies, stabilization strategies, structure-performance relationships, and experimental and theoretical advances in photocatalytic water splitting, CO2 reduction, biomass conversion, and related energy and small-molecule transformations. Finally, current challenges and prospects of atomically dispersed cocatalysts are discussed.

Communication
Visible-light photocatalytic 1,1-difunctionalizing carboxylation of isocyanides with CO2
Yuan-Xu Jiang, Zhi-Xin Yan, Long-Fu Wang, Yi-Chi Chen, Wen-Hao Xu, Shu-Rong Zhang, Jun-Ping Yue, Min Pan, Yong-Yuan Gui, Lei Song, Li-Li Liao, Jian-Heng Ye, Da-Gang Yu
2026, 91:  93-102.  DOI: 10.1016/S1872-2067(26)65211-4
Abstract ( 56 )   HTML ( 2 )   PDF (2122KB) ( 18 )  

Difunctionalizing carboxylation of unsaturated compounds with CO2 has emerged as a powerful strategy for synthesizing functionalized carboxylic acids. While substantial progress has been made, existing approaches have predominantly focused on 1,2-difunctionalizing carboxylation processes. Herein, we report the first 1,1-difunctionalizing carboxylation of isocyanides with CO2, enabling the efficient construction of diverse N-heteroaromatic carboxylic acids under mild conditions. This visible-light photocatalytic protocol displays broad substrate scope, good functional group tolerance, ready scalability and facile late-stage derivatization. Mechanistic studies and density functional theory calculations reveal that the generated electron-donor-acceptor complex could facilitate the formation of isocyanide- or CO2-derived radical anions, according to their redox potentials and stability. These radical anion intermediates could undergo following carboxylation to afford diverse N-heteroaromatic carboxylic acids.

Articles
Harnessing electronic structure modulation of Cu2+-mediated layered double hydroxide towards selective C2 pathway in CO2 reduction and plastic upcycling
Jian Yiing Loh, Feng Ming Yap, Elton Song-Zhe Mah, Xianhai Zeng, Wee-Jun Ong
2026, 91:  103-121.  DOI: 10.1016/S1872-2067(26)65182-0
Abstract ( 81 )   HTML ( 3 )   PDF (10654KB) ( 38 )  
Supporting Information

The integration of CO2 electroreduction with plastic upcycling offers a sustainable route to generate high-value C2 products. This study presents a self-supported CuMgAl layered double hydroxide coupled with reduced graphene oxide (CMA-0.1-G) as a bifunctional electrocatalyst for both CO2 reduction (CO2RR) and polyethylene terephthalate (PET) oxidation (POR). The hierarchical 1D/3D nanoneedle structure enables abundant active sites, efficient charge transport, and high stability. CMA-0.1-G achieves a 51.98% Faradaic efficiency (FE) for C2 products during CO2RR, attributed to mixed Cu2+/Cu+/Cu0 oxidation states that promote *CO adsorption and C-C coupling. In-situ Raman spectroscopy confirms *CO intermediates and strong Cu-CO interactions. For POR, the catalyst demonstrates an impressive > 89% FE toward C2 oxidation products, aided by redox-active M-OOH species. A two-electrode bifunctional system using CMA-0.1-G for both CO2RR and POR is assembled and operates at 1.25 V for 10 mA cm-2. In addition, the catalysts were tested using flow cell, demonstrating impressive performance for over 12 h under high voltage. Density functional theory analysis reveals the key intermediates formation, adsorption and desorption of the intermediates on key catalytic active sites, offering mechanistic insights into the optimum catalyst. Under simulated sunlight, the system achieves 5.68% solar-to-fuel efficiency for CO2RR and 92% FE for oxalic acid. This work establishes a multifunctional, scalable platform for coupling CO2RR and plastic upcycling, selectively generating C2 products through coordinated redox reactions, offering a viable solution for carbon neutrality and circular chemical manufacturing.

Zn-mediated modulation of Fe-Si interactions to achieve a balance between stability and selectivity in CO2 hydrogenation
Yi Liu, Peng Zhang, Yijun Li, Zhihua Han, Bo Feng, Xiaowa Nie, Jingyun Chen, Ruonan He, Zhoujun Wang, Lei Shao, Shaodong Qin, Guanghui Zhang, Xinwen Guo
2026, 91:  122-134.  DOI: 10.1016/S1872-2067(26)65216-3
Abstract ( 59 )   HTML ( 3 )   PDF (16908KB) ( 11 )  
Supporting Information

The direct hydrogenation of CO2 to high-value olefins represents one of the most attractive pathways for large-scale CO2 conversion. However, developing catalysts that are active, selective, and stable for olefin production remains challenging, particularly when incorporating support or binder under industrially relevant conditions. Although silica (SiO2) is widely used as a support or binder in industrial Fe-based Fischer-Tropsch synthesis (FTS), its incorporation often leads to poor performance in CO2 hydrogenation. In this work, we introduce a Zn promoter to modulate the detrimental Fe-Si interaction in SiO2-containing Fe-based catalysts for CO2 hydrogenation. The resulting catalyst achieves a single-pass C2+ olefin yield as high as 0.6 g·gcat-1·h-1, coupled with exceptional operational stability for over 500 h at high space velocity. Multiple characterization techniques reveal that during the in-situ reaction-activation stage, Zn promotes the formation of a Zn2SiO4 phase, which alleviates the adverse Fe-O-Si interaction. The high content of χ-Fe5C2 with electron-enriched surface and abundant surface oxygen vacancies contribute to the outstanding catalytic activity and selectivity, while the presence of Zn2SiO4 contributes to long-term stability. These findings provide valuable insights for the rational regulation of metal-support/binder interactions in CO2 hydrogenation catalytic systems, especially for the design of catalysts where shaping processes and long-term durability pose significant challenges.

Efficient urea synthesis via electrocatalytic CO2 reduction reaction coupled with nucleophilic addition reaction of NH3
Shuchang Fan, Peize Li, Tianyu Sun, Yan Wei, Zhaoshi Yu, Jiandong Zhang, Mingkui Wang, Yan Shen
2026, 91:  135-146.  DOI: 10.1016/S1872-2067(26)65213-8
Abstract ( 56 )   HTML ( 2 )   PDF (5200KB) ( 9 )  
Supporting Information

The electrocatalytic CO2 reduction reaction coupled with nucleophilic addition reaction of NH3 represents a promising route to synthesis urea. Relying on interfacial reactions between surface-adsorbed CO2•- from CO2 reduction and dissolved NH3. However, its efficiency is limited by competing CO formation and weak NH3 adsorption, which restrict the C-N coupling efficiency. Thus, achieving synergistic co-adsorption of CO2•- and NH3 while suppressing the CO pathway is essential. This study presents a dual-site catalyst with Pd clusters and Ni single atoms (Pd1.0/NiSAs-N-C). In 0.5 mol·L-1 NH4HCO3 solution at -1.3 V vs. RHE, Pd1.0/NiSAs-N-C achieves a record urea production rate of 3.10 mmol·L-1·g-1·h-1, substantially exceeding NiSAs-N-C (0.22 mmol·L-1·g-1·h-1). Experimental and theoretical analyses reveal that electronic interaction between Ni and Pd sites induces charge redistribution, which enhances NH3 adsorption on Pd via d-p orbital hybridization and optimizes CO2•- binding on Ni. This synergy suppresses CO formation and promotes CO2•- transfer to Pd sites, thereby increasing the “collision” probability between CO2•- and NH3 and reducing the C-N coupling energy barrier from 0.40 to 0.24 eV, which accelerates the overall kinetics. The “site division-intermediate migration” mechanism offers a new paradigm for electrocatalytic coupling systems.

Unraveling the surface structural evolution and single-atom doping effect in CO2-assisted ethane dehydrogenation over TiO2-based catalysts
Kexin Ma, Mifeng Xue, Yuhan Wang, Maohong Fan, Lixia Ling, Riguang Zhang
2026, 91:  147-162.  DOI: 10.1016/S1872-2067(26)65205-9
Abstract ( 48 )   HTML ( 3 )   PDF (5747KB) ( 7 )  
Supporting Information

CO2-assisted ethane dehydrogenation (CO2-ODHE) provides a promising route for ethylene production; however, the dynamic evolution of catalyst surface and its influence on the reaction mechanism are still not fully elucidated. In this study, we combined density functional theory calculations and kinetic Monte Carlo simulations to explore the surface evolution and catalytic behavior of CO2-ODHE over TiO2-based catalysts. The results show that the pristine TiO2 follows the Mars-van Krevelen (MvK) mechanism, where CO2 replenishes oxygen vacancies, but exhibits limited activity. Doping with single-atom metal (M-TiO2-Ov) suppresses CO2 dissociation at oxygen vacancy sites, shifting the reaction mechanism from MvK to a coupling mechanism that combines ethane dehydrogenation with the reverse water-gas shift reaction. In this coupled mechanism, CO2 primarily participates in hydrogenation via the formation of a COOH* intermediate, thereby efficiently removing surface hydrogen species. Among 15 types of M-doped TiO2 catalysts, Pd-TiO2 catalyst demonstrates the highest C2H4(g) formation activity (TOF=4.62 × 10⁴ s-1) with 99.9% selectivity under optimal conditions (873 K, C2H6/CO2=2: 1), which is seven orders of magnitude higher than that of undoped TiO2. Furthermore, the catalytic activity exhibits a linear correlation with the adsorption energy of H*, identifying the surface hydrogen adsorption strength as an effective descriptor for predicting C2H4(g) formation activity. This work provides a theoretical foundation and structural insights for designing high-performance CO2-ODHE catalysts via second-metal doping to regulate the reaction mechanism and catalytic performance of metal oxide.

Machine learning-accelerated electrocatalytic microkinetic model for global reaction network simulation of CO2 reduction reaction
Xiao-Li Liang, Zhao-Yu Wang, Shi-Lin Yan, Ting-Ting Yin, Wei Zhang, Hao Zhang, Xiaohong Liu, Liu-Bin Zhao
2026, 91:  163-175.  DOI: 10.1016/S1872-2067(26)65208-4
Abstract ( 15 )   HTML ( 4 )   PDF (4644KB) ( 4 )  
Supporting Information

Electrochemical CO2 reduction represents a sustainable route for energy storage and carbon utilization. Despite copper-based catalysts uniquely enabling hydrocarbons and oxygenates product formation, advancing this technology requires deeper mechanistic insights and accelerated catalyst screening. Herein, we establish a novel electrocatalysis research framework integrating three synergistic components: (1) With a force convergence accuracy of 0.02 eV/Å, the AdsorbML-FT workflow can achieve a 24-fold acceleration in DFT optimization steps whilst accounting for adsorption site screening, thereby enabling more efficient energy calculations for general substrate-adsorbate systems; (2) the global reaction network simulation of CO2RR enabling broader coverage of reaction mechanism including 22 surface species and 40 elementary reactions beyond traditional optimal-path limitations; (3) the electrocatalytic microkinetic model ensuring greater accuracy in predicting both catalytic activity and product selectivity. This integrated approach quantitatively decodes the CO2-to-C1 mechanism on Cu(111) and systematically screens 37 Cu3M1 alloys. We identify Cu3Al1 as an optimal catalyst, driving CO conversion at relatively low overpotentials with simultaneously high catalytic activity and CH4 product selectivity. The proposed machine learning-accelerated electrocatalytic microkinetic model bridges theoretical simulations with experimental measurements, offering a transformative tool for rapid electrocatalyst discovery.

Low CeO2 content with superior performance: In-situ spectroscopic insights into multicomponent CuFeCe/γ-Al2O3 catalysts for mild-temperature reverse water-gas shift
Deng Min, Sebastian Cisneros, Abdallah I. M. Rabee, Thanh Huyen Vuong, Jiadong Xiao, Yaoyuan Zhang, Rongsheng Cai, Feng Shi, Jabor Rabeah
2026, 91:  176-191.  DOI: 10.1016/S1872-2067(26)65221-7
Abstract ( 40 )   HTML ( 2 )   PDF (5248KB) ( 12 )  
Supporting Information

CO is a key precursor for high-value-added chemicals, and its production via the reverse water-gas shift reaction is crucial for carbon-neutral processes. Herein, we report on a multicomponent low Ce-containing CuFeCe/γ-Al2O3 catalyst (actual loading Cu=0.84 wt%, Fe=0.46 wt% and Ce=4.5 wt%) which achieves CO formation rate of ca. 47 mmol gcat-1 h-1 with > 99% selectivity at 380 °C, surpassing the performance of bulk CeO2-based catalysts under similar conditions. Using in-situ diffuse reflectance infrared Fourier transform spectroscopy, in-situ electron paramagnetic resonance and ex-situ X-ray photoelectron spectroscopy, we demonstrate that the superior activity arises from the coexistence of Cu single sites and small Cu0 clusters, stabilized by FeOx clusters and neighboring O-vacancies (Ov) associated with highly dispersed CeOx nanoparticles on γ-Al2O3. Cu single sites vicinal to Ce3+ rapidly shuttle between Cu2+ and Cu+, activating CO2 while Cu0 clusters facilitate H2 dissociation. In addition to direct CO2 activation, H-assistance pathway via a bridged formate intermediate contributes to CO formation, preferentially at the interfaces shared by Cu, Fe, and Ce species. Compared to monometallic and bimetallic analogs, the trimetallic catalyst exhibits markedly higher activity. Long-term tests (115 h) reveal reversible re-activation cycles, demonstrating the catalyst’s robustness under dynamic operating conditions.

Phosphorus vacancy-engineered Ru,Pv-CoP as a hydrogen spillover catalyst for enhanced industrial alkaline water electrolysis
Juan Xiao, Tingting Huang, Xia Liu, Yuying Feng, Guancheng Xu, Li Zhang
2026, 91:  192-209.  DOI: 10.1016/S1872-2067(26)65195-9
Abstract ( 63 )   HTML ( 2 )   PDF (7896KB) ( 7 )  
Supporting Information

Developing industrial-grade electrocatalysts based on the hydrogen spillover (HSo) mechanism is a key approach for translating lab-scale performance into industrial applications. Herein, a novel HSo catalyst (Ru,Pv-CoP/CF) was constructed by introducing phosphorus vacancies (Pv) into a CoP support decorated with Ru nanocrystals (Ru NCs) and Ru single atoms (Ru SAs) via defect-engineering strategy. The creation of Pv modulates the interfacial properties between Ru species and CoP, enabling electronic coupling between Ru and Co through phosphorus bridging and forming a “Ru-Co” donor-acceptor electron pair. Multidimensional in-situ electrochemical measurements reveal that Pv accelerates the polarization/dissociation of water molecules and the capture/migration of adsorbed hydrogen (H*). Further density functional theory calculations underscore that Pv narrows the work function difference between Ru NC and RuSA-CoP, which enhances the electron supply from Ru NC to Co, thereby triggering an efficient H* cross-phase transmission. Specifically, Ru NCs primarily facilitate H2O dissociation, and the generated H* subsequently spillover to adjacent Co sites for H2 desorption. Benefiting from the Pv-mediated relay catalysis, the Ru,Pv-CoP/CF catalyst exhibits top-level HER properties in 1.0 mol L-1 KOH, requiring only 86 mV at an industrial current density of 1.0 A cm-2 with exceptional durability over 1200 h. This work advances the fundamental understanding of vacancy-guided HSo mechanism, providing unique insights for designing transition metal phosphides-based electrocatalysts for industrial alkaline water electrolysis.

Activating synergetic adsorbate evolution and lattice oxygen mechanisms in NiFe-LDH via M-O-Si bonding for improved oxygen evolution reaction
Jing Xie, Ying Tang, Songbo Ye, Wentao Bao, Tiantian Wang, Yunxia Zhao, Shifu Wang, Jie Yu, Xuning Li, Hao Li, Feng Yu
2026, 91:  210-221.  DOI: 10.1016/S1872-2067(26)65225-4
Abstract ( 64 )   HTML ( 2 )   PDF (6885KB) ( 11 )  
Supporting Information

High overpotential oxygen evolution reactions (OER) remain a strongly challenging factor for efficient hydrogen production via water electrolysis in energy harvesting devices. Here, we propose a new strategy for lowering overpotentials based on M-O-Si (M=Ni, Fe) covalent bond formation through a rapid 30-s chemical etching process to create a silicon-doped anemone-like NiFe layered double hydroxide (Si-NiFe-LDH/NF). Our strategy decreases the catalytic phase-transition potential and optimizes intermediate binding at Ni sites, reducing the adsorbate evolution mechanism barrier. Concurrently, it induces an upshift of the O 2p band center, which activates lattice oxygen and lowers the energy barrier along the lattice oxygen mechanism pathway. Our findings highlight the synergistic optimization of the dual mechanisms, demonstrating superior catalytic performance of Si-NiFe-LDH/NF in 1 mol L-1 KOH relative to NiFe-LDH/NF, where OER overpotentials are reduced by 24% and 14.4% at current densities of 10 and 100 mA cm-2, respectively. In addition, a low hydrogen evolution reaction overpotential of 112 mV at 10 mA cm-2 is achieved. Finally, we demonstrate the strong potential of our strategy in industrial applications through the operation (at 1.55 V) of an assembled solar-powered total water electrolyzer.

Interfacial engineering of an advanced NiV-LDH@MoS2 heterostructure electrocatalyst for efficient alkaline hydrogen evolution
Shihao Yu, Yueying Yan, Yang Yang, Bohan Yao, Ziwen Zhu, Dewen Wang, Zhicai Xing, Xiurong Yang
2026, 91:  222-231.  DOI: 10.1016/S1872-2067(26)65224-2
Abstract ( 53 )   HTML ( 2 )   PDF (1977KB) ( 22 )  
Supporting Information

The development of efficient and stable non-noble-metal electrocatalysts is crucial for alkaline water electrolysis toward hydrogen production. Herein, a heterostructure comprising NiV-LDH decorated MoS2 nanosheet arrays on carbon cloth (NiV-LDH@MoS2) is fabricated via a facile two-step hydrothermal method. The catalyst exhibits outstanding hydrogen evolution reaction (HER) performance in 1 mol L-1 KOH, requiring an overpotential of only 103 mV to deliver a current density of 10 mA cm-2, with a Tafel slope of 76.9 mV dec-1 and exceptional stability surpassing 1000 h. The characterizations combined with theoretical calculations reveal that the heterointerface induces electron transfer from NiV-LDH to MoS2, effectively modulating the electronic structure. This synergy significantly lowers the water-dissociation energy barrier and optimizes the hydrogen adsorption free energy, thereby collectively accelerating the alkaline HER kinetics. Furthermore, an anion-exchange membrane water electrolyzer assembled with this catalyst achieves a high current density of 1 A cm-2 at 2.26 V and maintains stable operation for over 200 h, underscoring its practical potential. This work provides an effective interfacial-engineering strategy for designing high-performance, low-cost electrocatalysts for the alkaline HER.

Chromium-doped CoFe2O4 for enhanced oxygen evolution via hydroxyl spillover
Muhammad Umair Ul Hassan, Jizhang Wang, Rashid Mehmood, Fazal Shah, Kun Qi, Fuxiang Zhang
2026, 91:  232-240.  DOI: 10.1016/S1872-2067(26)65231-X
Abstract ( 60 )   HTML ( 2 )   PDF (15722KB) ( 31 )  
Supporting Information

Developing efficient, low-cost oxygen evolution reaction (OER) electrocatalysts capable of achieving high current densities is critical for advancing practical water electrolysis. Herein, we report a chromium-doped cobalt ferrite (i.e., Cr-CoFe2O4/IF) as an efficient OER electrocatalyst in alkaline media. The Cr-CoFe2O4/IF electrode exhibits overpotentials of 260 and 300 mV at current densities of 100 and 500 mA cm-2, respectively, in a three-electrode system and maintains stable operation for more than 400 h, demonstrating its potential for practical application. Structural characterization demonstrates that Cr doping modulates the electronic structure of the Co/Fe active centers. Density functional theory calculations reveal that Cr incorporation introduces preferential hydroxyl adsorption sites, thereby decreasing the Gibbs free energy of the OER process. In particular, *OH species adsorbed on Cr sites can migrate to adjacent Co/Fe active sites through a dopant-mediated hydroxyl spillover pathway, which facilitates the formation of O-O coupling to lower the reaction energy barrier. This work establishes dopant-mediated hydroxyl spillover as an effective strategy for designing earth-abundant spinel oxides toward high-rate OER electrocatalysis.

Boosting sequential single-electron O2 reduction reaction on S-scheme SbVO4/PCN heterojunction for efficient artificial photosynthesis of H2O2 under ambient condition
Zhuangzhuang Wang, Yuan Li, Qiang Cheng, Xiaotian Wang, Bin Liu, Gaoke Zhang
2026, 91:  241-251.  DOI: 10.1016/S1872-2067(26)65203-5
Abstract ( 66 )   HTML ( 2 )   PDF (3758KB) ( 10 )  
Supporting Information

Non-sacrificial artificial photosynthesis of H2O2 offers a charming process for onsite H2O2 generation with unique merits of sustainability and environmental friendliness. However, the low conversion efficiency of sequential single-electron oxygen reduction reaction (SSORR) greatly limits the H2O2 production rate. Herein, we develop a step-scheme (S-scheme) SbVO4/PCN system for artificial photosynthesis of H2O2, which displays greatly boosted photocatalytic SSORR. The H2O2 yield can reach as high as 94.7 μmol·g-catalyst-1·h-1 under visible light illumination (λ > 420 nm), about 41 times higher than that of the pristine PCN. The S-scheme charge transfer pathway is systematically demonstrated by photo-irradiated Kelvin probe force microscopy, in-situ X-ray photoelectron spectroscopy and surface photovoltage measurements. Theoretical calculations further indicate that the SbVO4/PCN is favorable to transfer electrons from its hybrid p orbitals to the 1π and 2π* orbitals of O2 and the evolution from *O2 to *OOH consumes no energy, therefore greatly promoting the formation of PCN-*OOH even without electron donor. The onsite generated O2 via water oxidation reaction on SbVO4 can replenish its consumption on PCN, leading to boosted reaction kinetics.

Synergy of spatially adjacent redox sites and spin polarization in ZnIn2S4 for boosted H2O2 photosynthesis
Zhiqi Li, Chen Guan, Kaiqiang Xu, Wei Xia, Dilshod D. Nematov, Quanjun Xiang
2026, 91:  252-262.  DOI: 10.1016/S1872-2067(26)65206-0
Abstract ( 70 )   HTML ( 2 )   PDF (5948KB) ( 17 )  
Supporting Information

Proton-coupled electron transfer (PCET) is essential for driving the two-electron oxygen reduction pathway toward H2O2 production. However, the coupled integration of the adjacent water oxidation reaction and oxygen reduction reaction sites to amplify PCET presents a considerable challenge in the photocatalytic H2O2 production. Addressing this challenge requires overcoming the inevitable carrier recombination problem at spatially adjacent redox sites. In this work, we incorporated Co and sulfur vacancies into ZnIn2S4 as a photocatalyst, in which the incorporation of Co induced spin polarization, significantly inhibiting the recombination of charge carriers. While the sulfur vacancies created an unsaturated coordination environment at the metal sites to promote the adsorption of reactants, forming the spatially adjacent redox sites. Extended X-ray absorption fine structure results and density functional theory simulations revealed the formation of Co-Vs dual sites. In-situ Fourier transform infrared and Raman spectroscopy confirmed the promoted PCET. This resulted in a 3.6-fold increase (215.4 μmol g-1 h-1) in H2O2 yield compared to pure ZIS (59.6 μmol g-1 h-1). This work exhibited a strategy for constructing spatially synergistic sites and manipulating the electronic spin polarization, providing an effective approach to enhance photocatalytic H2O2 production.

Hole-confining covalent organic frameworks for photosynthesis of H2O2 and H2 from pure water
Zeng Li, Mingfei Yu, Liuyi Li, Yan Yu
2026, 91:  263-273.  DOI: 10.1016/S1872-2067(26)65148-0
Abstract ( 5 )   HTML ( 2 )   PDF (8092KB) ( 6 )  
Supporting Information

Covalent organic frameworks (COFs) have emerged as promising candidates for photocatalysis, however, their efficiency in converting water into value-added chemicals is often severely constrained by sluggish water-oxidation kinetics. Herein, we demonstrate a strategy of confining photogenerated holes at specific water oxidation active sites in COFs to enhance simultaneous H2 and H2O2 co-generation from water. We designed three hydrazone-linked COFs capable of driving overall water splitting without the use of sacrificial agents. Mechanistic studies reveal that the capability to confine photogenerated holes at hydrazone linkages directly correlates with superior water-splitting activity. Specifically, the accumulation of holes at these thermodynamically preferred water-adsorption sites, coupled with electron delocalization, results in a unique spatial charge separation. This configuration not only significantly suppresses recombination to extend excited-state lifetimes but also creates hole-rich local environments that accelerate water oxidation kinetics. Consequently, the optimal COF achieves H2 and H2O2 production rates of 311.2 and 278.6 μmol·g-1·h-1, respectively. This work establishes the spatial accumulation of photogenerated holes as a pivotal, engineerable design principle for advancing COF-based photocatalytic water splitting.

Exploring the spatially resolved charge transfer mechanism in the Al-doped SrTiO3/C3N5 S-scheme heterojunction for boosted solar-driven photocatalytic hydrogen evolution
Xiaoxuan Chen, Kang Li, Zewen Jiang, Haotian Guo, Xuanwen Xu, Pengyu Dong
2026, 91:  274-289.  DOI: 10.1016/S1872-2067(26)65226-6
Abstract ( 64 )   HTML ( 2 )   PDF (11790KB) ( 17 )  
Supporting Information

Developing highly efficient and low-cost C3N5-based photocatalysts for hydrogen production is of great significance. However, research on the surface potential distribution of photogenerated charges in the C3N5-based S-scheme heterojunction from a spatial perspective with nanoscale precision is still lacking. In this work, using Al-doped SrTiO3 (ASTO) with strong oxidation ability and C3N5 nanosheets with high reduction ability, a novel S-scheme heterojunction was created. The ASTO/C3N5 S-scheme heterojunction was synthesized via a hydrothermal method. It reveals that the polyhedral ASTO particles are in intimate contact with C3N5 nanosheets, with a clear and distinct interface observed between them. The optimal 5%-ASTO/C3N5 S-scheme heterojunction exhibits a high hydrogen production rate under simulated solar light, surpassing pristine C3N5 and ASTO by factors of 2.5 and 37.4, respectively. Light-assisted Kelvin-probe force microscopy was employed to uncover the spatially resolved features of the kinetics of charge transfer in the ASTO/C3N5 S-scheme heterojunction. It reveals that photogenerated electrons gather on the surface of C3N5 as a reduction photocatalyst in the heterojunction, resulting in a reduced surface photovoltage (SPV) compared to pure C3N5, whereas holes accumulate on the surface of ASTO as an oxidation photocatalyst in the heterojunction, providing higher SPV than pure ASTO. Moreover, the large work function difference (0.41 eV) between ASTO and C3N5 induces a robust interfacial electric field (IEF). This strong IEF facilitates the photogenerated electrons in ASTO’s conduction band recombine with holes in C3N5’s valence band, preserving high redox potentials. Furthermore, active free radical measurement confirms the enhanced •O2- and •OH signals for the 5%-ASTO/C3N5 heterojunction, confirming the S-scheme mechanism and excluding the type-II mechanism, which is also demonstrated by in-situ X-ray photoelectron spectroscopy. Overall, this work offers some evidence regarding the spatially resolved characteristics of the kinetics of charge transfer in the S-scheme heterojunction.

Engineering chemically bonded S-scheme heterojunction between 5,10,15,20-tetrakis(4-sulfophenyl)porphyrin and Bi4O5Br2 for efficient photocatalytic water purification
Shijie Li, Yanbo Wang, Kexin Dong, Fang Yang, Yang Guo, Xiaobing Wang, Yanping Hou, Bin Zhang
2026, 91:  290-303.  DOI: 10.1016/S1872-2067(26)65237-0
Abstract ( 88 )   HTML ( 2 )   PDF (5999KB) ( 31 )  
Supporting Information

The rational design of organic-inorganic heterojunction systems represents a promising strategy for developing robust photocatalysts in environmental remediation. Thus, this study pioneers a novel S-scheme heterojunction by integrating oxidative Bi4O5Br2 (BiBr) nanosheets with reductive tetraphenylporphyrin tetrasulfonic acid (TPPS) via Bi-O-S bonds, leveraging their intrinsically staggered band alignment. The optimized TPPS/BiBr heterojunction achieves a remarkable tetracycline degradation rate constant of 0.0265 min‒1 under visible light, 1.2-fold higher than pristine BiBr. This enhancement originates from a robust internal electric field that drives a directional S-scheme charge-transfer model. This unique mechanism not only propels electron migration from BiBr to TPPS via a well-established Bi-O-S atomic-scale transport channel but also strategically preserves the potent reduction capacity of the electrons accumulated in the TPPS lowest unoccupied molecular orbital, while simultaneously maintaining the strong oxidation power of the holes retained in the BiBr valence band. Radical quenching experiments and mechanistic analyses further corroborate that •O2-, •OH, and h+ drive significantly augmented antibiotic degradation performance. This work establishes a groundbreaking approach for constructing tailored chemically bonded organic-inorganic S-scheme photocatalysts, demonstrating superior interfacial charge dynamics while maximizing redox potential utilization for efficient antibiotic wastewater decontamination.

Interfacial charge transfer via SMSI in heterostructured Pd/Bi2O3: A paradigm for sustainable electrochemical biomass upcycling
Han-Yue Yang, Jia-Jia Wen, Xue Xiao, Yu Chen
2026, 91:  304-314.  DOI: 10.1016/S1872-2067(26)65201-1
Abstract ( 59 )   HTML ( 2 )   PDF (2174KB) ( 16 )  
Supporting Information

The electrochemical oxidation of ethylene glycol (EG) to glycolic acid (GA) represents a promising route for valorizing plastic waste and biomass-derived feedstocks while co-producing hydrogen. However, existing catalysts are limited by low selectivity, rapid deactivation, and high overpotentials. Herein, we report a 0D/2D heterostructured catalyst, Pd-NCs/Bi2O3-NSs, in which Pd nanocrystals are uniformly anchored on ultrathin Bi2O3 nanosheets. Spectroscopic studies and density functional theory calculations reveal that the strong Pd-O-Bi interfacial interactions induce charge transfer from Bi2O3 to Pd, generating electron-rich Pd sites with a downshifted d-band center. Together with the effect of oxophilic Bi2O3, this optimizes reaction energetics and suppresses intermediate poisoning. As a result, the catalyst delivers 5.81 A mgPd-1 mass activity, 92.1% Faradaic efficiency for GA, and excellent durability. Techno-economic and life-cycle analyses further confirm its cost competitiveness and environmental benefits.

Electrocatalysts based on oxygen vacancy-induced frustrated Lewis pairs for selective electrooxidation of ethylene glycol to formic acid
Xuanrui Yang, Wenxuan Huang, Jianhua Shen, Chunzhong Li, Yihua Zhu
2026, 91:  315-326.  DOI: 10.1016/S1872-2067(26)65200-X
Abstract ( 58 )   HTML ( 2 )   PDF (2434KB) ( 10 )  
Supporting Information

The sluggish oxygen evolution reaction (OER) limits alkaline water electrolysis efficiency, and replacing OER with ethylene glycol oxidation reaction (EGOR) is a promising strategy to reduce energy consumption while producing value-added chemicals. EGOR for formate/formic acid (FA) requires overcoming C-C bond breakage barriers, achievable via constructing frustrated Lewis pairs (FLPs) and tuning electron distribution. Herein, a series of layered double hydroxide (LDH) electrocatalysts were fabricated via hydrothermal-assisted self-etching assembly using Co/Ni foam as substrates. Ni-doped CoNiFe-LDH/CoFoam exhibited excellent EGOR performance, with a low onset potential, 1.37 V vs. RHE at 100 mA cm-2, and 88% Faradaic efficiency (FE) for FA. The in-situ characterizations showed Co sites lower onset potential, while Ni doping modulates electronic structure and induces oxygen vacancies (Ov). The theoretical calculations confirmed Ov forms surface FLPs, facilitating C-C bond cleavage and enhancing activity/selectivity. Notably, the catalyst performs well in polyethylene terephthalate (PET) degradation, enabling simultaneous plastic valorization and low-energy hydrogen production, providing insights for designing high-performance electrocatalysts for sustainable energy and environmental applications.

Efficient and stable conversion of ethanol/acetaldehyde to 1,3-butadiene over a Zr1/NC@SBA-15 single-atom catalyst
Yujia Zhao, Xianquan Li, Rizheng Jing, Jifeng Pang, Wenguang Yu, Hua Wang, Feifei Xu, Mingyuan Zheng
2026, 91:  327-340.  DOI: 10.1016/S1872-2067(26)65130-3
Abstract ( 103 )   HTML ( 2 )   PDF (6828KB) ( 22 )  
Supporting Information

The design of high-performance single-atom catalysts (SACs) with excellent stability for the ethanol-to-1,3-butadiene (1,3-BD) (ETB) process remains a significant challenge and has rarely been reported. Herein, a Zr single-atom catalyst Zr1/NC@SBA-15 was constructed and employed in the ETB process. Compared with the conventional nano-Zr/SBA-15 catalyst, the Zr1/NC@SBA-15 catalyst delivered notably higher performance: the reactant conversion exceeded 80%, the selectivity toward 1,3-BD was above 70%, and unprecedentedly high stability of 300 h on stream was achieved without notable deactivation. Systematic characterizations, operando spectroscopy, and density functional theory calculations revealed that the Zr atoms were uniformly dispersed on the Zr1/NC@SBA-15 catalyst in the single-atom state, forming abundant Lewis acid sites (LAS) of [(Si-N)-Zr(O-Si)2(OH)]. A dual acetaldehyde adsorption mode was proposed for the ETB reaction over the Zr1/NC@SBA-15 catalyst. The rate-determining step of acetaldehyde condensation proceeded via the Langmuir-Hinshelwood mechanism over the [(Si-N)-Zr(O-Si)2(OH)] sites, with apparent activation energy decreased by 18.8 kJ/mol compared with that over the nano-Zr/SBA-15 catalyst. The N-coordinated single Zr atom with lowered energy endows the Zr1/NC@SBA-15 catalyst with outstanding stability during the reaction; meanwhile, the unique coordination of the [(Si-N)-Zr(O-Si)2(OH)] sites and the enhanced LAS acidity result in the high activity of Zr1/NC@SBA-15 in the ETB reaction. This work provides a good example of SAC application in the ETB process and opens a new route for exploring more effective catalysts.

Phosphate anions accelerated deprotonation and proton transfer for water-involved electrooxidation
Junwei Yao, Chuanqi Cheng, Rui Li, Ying Gao, Bin Zhang
2026, 91:  341-350.  DOI: 10.1016/S1872-2067(26)65234-5
Abstract ( 86 )   HTML ( 3 )   PDF (5091KB) ( 35 )  
Supporting Information

Phosphate anions (Pi)-modified layered double hydroxides (LDH) have long been regarded as promising anodic electrocatalysts, but the effect by which Pi enhances electrooxidative activity remains unclear. Here, we prepare a Pi-intercalated NiFe LDH (Pi-NiFe LDH) to investigate the role of Pi intercalation in electrooxidation processes. Pi anchored at Fe sites facilitates the deprotonation of adjacent hydroxyl groups at Ni sites to form the active NiOOH phase while promoting the deprotonation of reactants to initiate electrooxidation processes. These dual effects collectively lower the thermodynamic energy. Moreover, Pi bridges the interfacial hydrogen-bond (H-bond) network to restore its continuity, ensuring efficient proton transfer and accelerating reaction kinetics. Therefore, Pi-NiFe LDH delivers enhanced activity for both the oxygen evolution reaction and the electrooxidation of various alcohols with reduced overpotentials and elevated yield rates. This study offers mechanistic insights into how anions modulate deprotonation and proton transfer processes, guiding the design of electrooxidation catalysts.

Highly stable encapsulated Ni catalyst for phenol hydrogenation: Synthesis and mechanistic insight
Maochen Qian, Wen Liu, Xinbao Zhang, Xiangxue Zhu, Xiujie Li
2026, 91:  351-364.  DOI: 10.1016/S1872-2067(26)65220-5
Abstract ( 58 )   HTML ( 3 )   PDF (6300KB) ( 20 )  
Supporting Information

Hydrogenation of biomass-derived phenol to cyclohexanol represents a sustainable pathway for cyclohexanol production. Although supported Ni catalysts exhibit potential for fixed-bed applications, traditional systems suffer from rapid deactivation. Herein, we demonstrate a highly stable encapsulated Ni@silicalite-1 catalyst synthesized via chelator-free hydrothermal route. Compared to conventional impregnated Ni/silicalite-1 counterparts, Ni@silicalite-1 maintains exceptional operational stability beyond 100 h with sustained phenol conversion exceeding 90% and cyclohexanol selectivity over 95% under continuous-flow conditions. Characterization results including X-ray absorption spectroscopy, in-situ diffuse reflectance infrared Fourier transform spectroscopy and high angle annular dark field scanning transmission electron microscopy reveal that ultrafine Ni nanoparticles (~2.34 nm) anchored via Ni-O-Si interfacial bonds constitute the active sites, conferring dual resistance to thermal sintering and carbon deposition under reaction conditions. Mechanistic studies tracking silicalite-1 crystallization demonstrate that amorphous nickel silicate precursors gradually transform into spatially confined Ni nanoparticles driven by the organic template. This work establishes a universal paradigm for constructing sintering-resistant metal-zeolite catalysts through nanoscale engineering of metal-support interfaces.

Coordination reconfigured Ni single-atoms for scaled photothermal COx free hydrogen production from ammonia
Yachuan Wang, Runan Lei, Wei Wang, Qixuan Wu, Kaiwen Kang, Dachao Yuan, Zhibo Zhang, Yuan Tang, Linjie Gao, Yaguang Li, Jinhua Ye
2026, 91:  365-375.  DOI: 10.1016/S1872-2067(26)65207-2
Abstract ( 90 )   HTML ( 2 )   PDF (7978KB) ( 28 )  
Supporting Information

Ammonia (NH3) decomposition to produce COx free hydrogen is an ideal supply for hydrogen refueling stations with the drawback of giant secondary energy consumption, which limits its application. Herein, we develop a natural sunlight driven photothermal NH3 decomposition system to achieve a hydrogen evolution rate of 298.0 mmol g-cat–1 h–1 with the solar-to-hydrogen efficiency of 16.1%, outperforming all counterparts. The catalytic activity is attributed to the catalyst of Ni single atoms loaded on a cerium-zirconium solid solution (SA Ni/CexZr1–xO2), that the component regulation of CexZr1–xO2 could modulate the Ni–O coordination number and shorten the Ni–O bond length of Ni single atoms to synergistically optimize the dissociative energy barrier of NH3 molecules and the desorption kinetics of NHx intermediates. Under outdoor sunlight illumination, the enlarged photothermal NH3 decomposition system achieves a record hydrogen production rate of 1 kg per day and the corresponding cost is calculated as $3.53 kg–1, significantly lower than the market price of hydrogen from thermocatalytic NH3 decomposition ($4.26 kg–1). This work provides a new strategy for NH3 decomposition in COx free hydrogen energy system with low energy consumption and cost advantage through the synergistic of coordination structure modulation of catalyst and photothermal system.

Enhanced Fe(III)/Fe(II) cycle in heterogeneous photocatalysis-self-Fenton system for organic pollutants degradation
Rong Ji, Jingran Yi, Xinyu Sun, Yuming Dong, Chengsi Pan, Hui Zhao, Yongfa Zhu
2026, 91:  376-385.  DOI: 10.1016/S1872-2067(26)65034-6
Abstract ( 49 )   HTML ( 3 )   PDF (6313KB) ( 13 )  
Supporting Information

Heterogeneous photocatalysis-self-Fenton systems have emerged as a promising method for wastewater treatment. However, their reaction efficiency is often severely limited by the sluggish regeneration of ferrous (Fe(II)). In this work, we report a novel iron(Fe)-based conjugated polymer capable of simultaneously achieving 2-electron photocatalytic oxygen activation and Fe(III) reduction, thereby enabling efficient degradation of organic pollutants. The underlying mechanism of the accelerated Fe(III)/Fe(II) cycle and enhanced degradation performance within this system is also elucidated. The polymer is constructed using a planar conjugated pyrene (Py) unit as the electron donor, which facilitates electron transfer for the reduction of Fe(III) to Fe(II) during H2O2 activation. Thiophene (Tp) and dibenzo[b,d]thiophene-s,s-dioxide (BDTO) are incorporated as the π-spacer and electron acceptor, respectively. The optimized Fe-Py-Tp-BDTO catalyst demonstrated outstanding performance, with an apparent rate constant of 0.14 min-1 for 4-chlorophenol degradation and a mineralization rate of 79.1%, which significantly surpasses those of current photocatalysts. Quenching experiments confirmed that hydroxyl radicals and holes are the predominant reactive species responsible for pollutant removal. This work successfully establishes a heterogeneous photocatalysis-self-Fenton system with a highly efficient Fe(III)/Fe(II) cycle, thereby broadening the practical application scope of conventional Fenton technology.

Preserving high reduction potential in 3D COFs through non-conjugated linkages for enhanced solar-driven O2 reduction
Zhongliang Wen, Maojun Deng, Xiao Feng, Pascal Van Der Voort
2026, 91:  386-395.  DOI: 10.1016/S1872-2067(26)65209-6
Abstract ( 60 )   HTML ( 4 )   PDF (12261KB) ( 37 )  
Supporting Information

Photocatalytic production of hydrogen peroxide (H2O2) via the two-electron oxygen reduction reaction (ORR) represents a sustainable alternative to the energy-intensive anthraquinone process. Covalent organic frameworks (COFs), with tunable π-conjugated structures and ordered porosity, have emerged as one of the most promising photocatalysts. However, the role of π-conjugation in governing photocatalytic activity remains poorly understood. Herein, we report a linkage-engineering strategy to systematically investigate the impact of conjugation on H2O2 production using a donor-acceptor (D-A) building unit, 2,3,5,6-tetrakis(3,6-bis(4-formylphenyl)-9H-carbazol- 9-yl)terephthalonitrile (TB). Three structurally analogous COFs with distinct linkage chemistries were constructed, including a conjugated imine-linked TB-DA COF, a weakly conjugated hydrazone-linked TB-TDA COF, and a non-conjugated urea-linked TB-PDU COF. Unexpectedly, photocatalytic evaluation under aqueous conditions reveals a reversed activity trend, with TB-PDU COF (1580 μmol g-1 h-1) outperforming TB-TDA COF (1288 μmol g-1 h-1) and TB-DA COF (523 μmol g-1 h-1). Mechanistic studies demonstrate that the non-conjugated urea linkage suppresses charge recombination, enhances hydrophilicity to facilitate O2 accessibility, and preserves favorable reduction potential by avoiding excessive electronic delocalization. These findings deviate from the conventional conjugation-driven design paradigm and highlight the critical role of linkage chemistry in balancing charge dynamics, interfacial properties, and reaction thermodynamics, providing new insights into the rational design of COF-based photocatalysts for efficient H2O2 production.

Balancing lattice oxygen activation and surface reoxidation in lanthanide-Zn co-doped Pd/CeO2 single-atom catalysts for CO oxidation
Mengrong Li, Bram D. Bruininks, Ivo A. W. Filot, Emiel J. M. Hensen
2026, 91:  396-411.  DOI: 10.1016/S1872-2067(26)65256-4
Abstract ( 11 )   HTML ( 5 )   PDF (4564KB) ( 7 )  
Supporting Information

Ceria-supported Pd single-atom catalysts are promising for low-temperature CO oxidation in three-way catalysis, yet the fundamental trade-off between stability and activity remains poorly understood. Here, we systematically investigate the effect of lanthanide (Ln)-Zn co-doping on Pd1/CeO2(111) using density functional theory and microkinetic simulations. In this design, Zn stabilizes isolated Pd atoms and promotes lattice-oxygen activation, while Ln dopants are introduced to further enhance lattice oxygen reactivity and tune reactant adsorption. Despite these intended benefits, microkinetic simulations reveal that Ln co-doping consistently decreases the CO oxidation activity compared to the Zn-doped reference. While Ln incorporation enhances lattice oxygen reactivity and facilitates vacancy formation, it simultaneously hinders O2 adsorption, activation, and vacancy replenishment. As a result, the catalytic cycle becomes limited by O2 activation rather than lattice oxygen participation, leading to a lower turnover frequency. These findings demonstrate that increasing lattice oxygen reactivity alone does not guarantee improved catalytic performance in systems operating via a Mars-van Krevelen mechanism. Instead, catalytic activity is governed by a delicate balance between lattice oxygen activation and reoxidation kinetics. Consequently, commonly used descriptors such as oxygen vacancy formation energy are insufficient to predict catalytic performance. This work highlights that effective catalyst design requires the selection of combined descriptors based on the kinetic characteristics of the specific catalytic system, rather than relying on a single universal descriptor.